Intact Mass Analysis Service
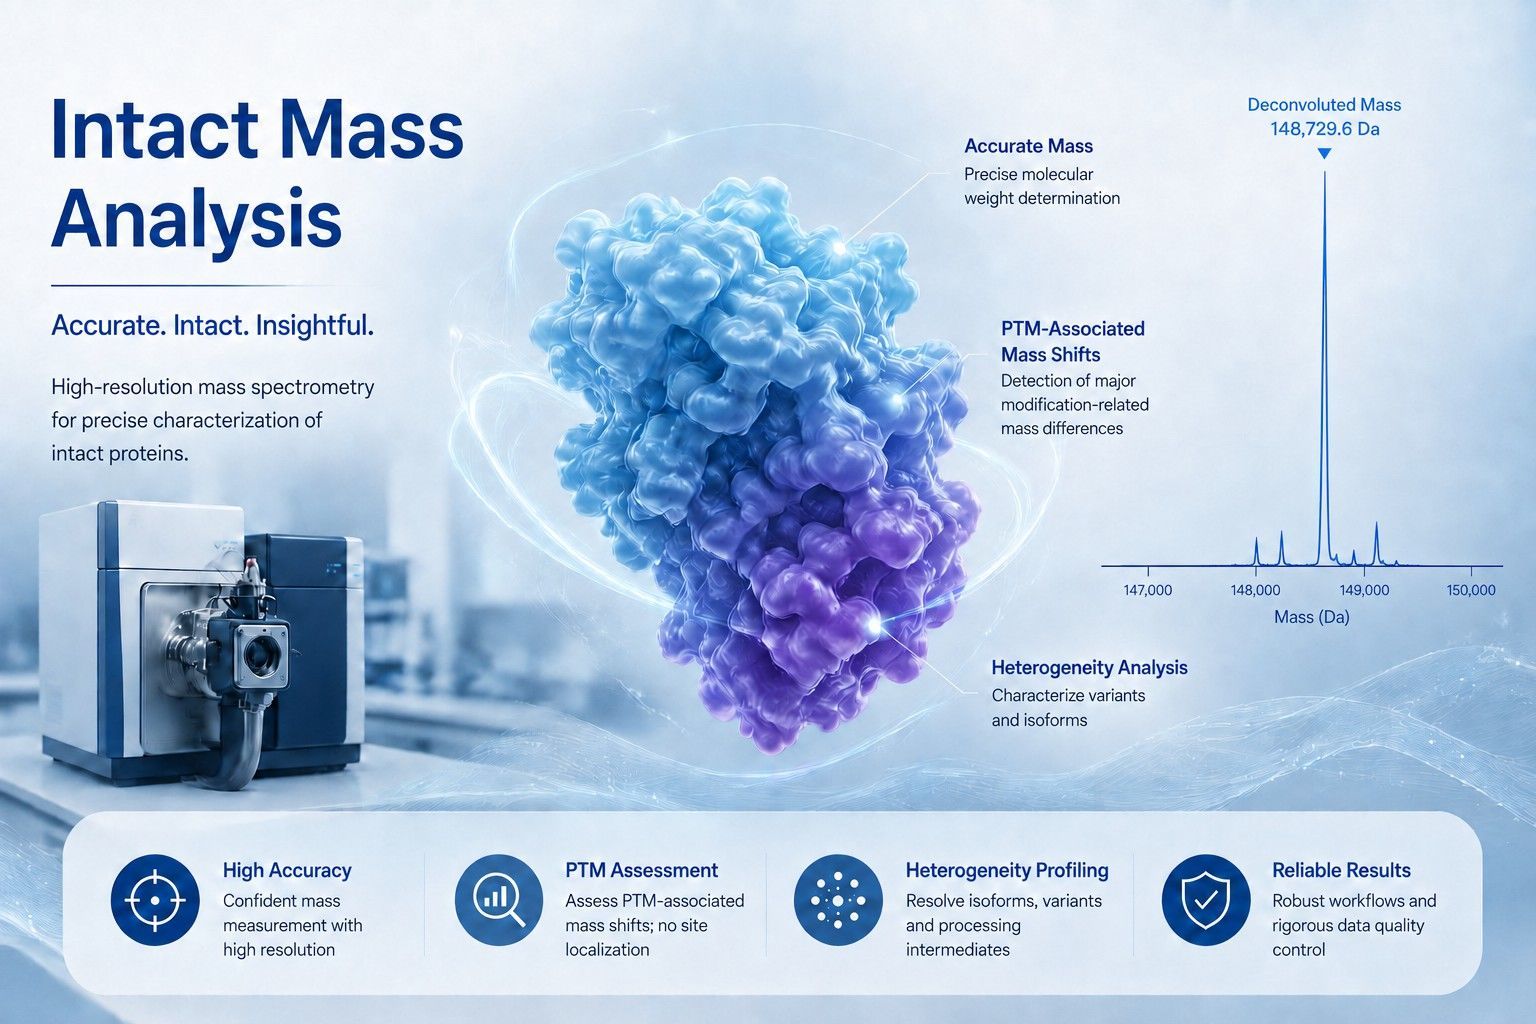
Our Intact Mass Analysis service provides high-resolution molecular weight determination for biotherapeutics, recombinant proteins, and other large molecules. Instead of breaking a protein into smaller fragments to study the pieces, we measure the entire, undigested molecule in its native or denatured state.
Designed for speed and precision, this approach delivers a direct, top-level view of your molecule. It can bypass lengthy digestion steps to support mass-based identity confirmation, reveal major mass shifts, and compare structural variants with confidence.
- Confirm observed molecular mass against the theoretical protein design.
- Screen for truncation, clipping, major modification-associated mass shifts, and unexpected attachments.
- Compare candidates, production batches, and reference materials efficiently.
In line with ICH Q6B Guidance for biologic characterization
Jump to Section
- Why Intact Mass Analysis Matters
- Service Capabilities
- Biologic Modalities
- Analytical Approaches
- Service Advantages
- Instrument Platform
- Workflow
- Report Deliverables
- FAQs
- Sample Preparation
Why Intact Mass Analysis Matters in Biologic Development
A biologic may contain a mixture of closely related molecular forms created by glycosylation, oxidation, terminal processing, clipping, incomplete assembly, or other manufacturing- and storage-related changes. Measuring the complete molecule provides a rapid way to confirm whether the observed mass agrees with the intended design and to screen for major variants that could affect downstream research decisions.
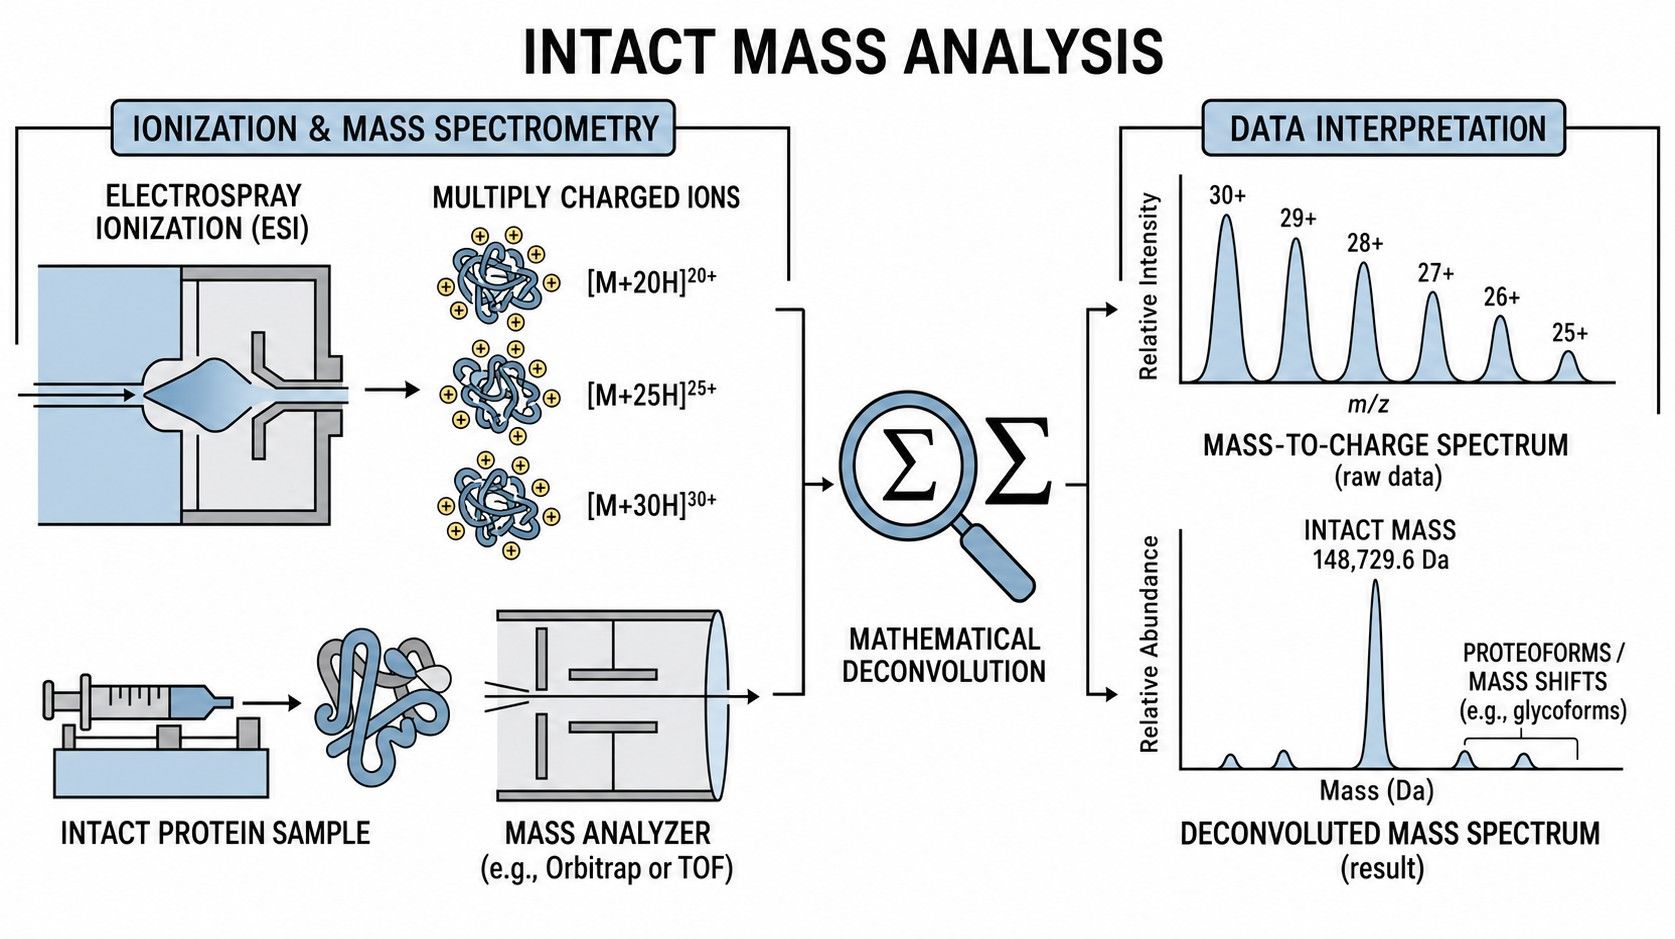
Intact Mass Analysis uses liquid chromatography coupled with high-resolution mass spectrometry to measure an entire, undigested protein. The resulting multi-charge spectrum is mathematically deconvoluted into a neutral molecular-mass profile, allowing the observed mass and major proteoforms to be compared with theoretical expectations.
Because the protein is evaluated without enzymatic digestion, intact analysis provides a fast whole-molecule overview with fewer preparation steps. It is valuable for identity confirmation, candidate screening, lot-to-lot comparison, degradation assessment, glycoform profiling, and antibody-drug conjugate characterization.
Intact mass analysis identifies global mass differences but generally does not localize a modification to a specific amino-acid residue. When site-level confirmation is needed, the results can guide orthogonal follow-up by reduced or subunit analysis, peptide mapping, glycan analysis, or targeted PTM characterization.
Intact Mass Analysis Service Capabilities
Our platform provides a direct whole-molecule view that supports the following core characterization needs:
Mass-Based Identity Confirmation
Compare the observed molecular weight with the theoretical design to support confirmation of the expressed protein or biotherapeutic.
Degradation and Clipping Detection
Screen for truncated proteins, chain loss, clipping, or other major mass changes that may indicate degradation or incomplete processing.
Global Proteoform Profiling
Reveal major molecular forms associated with glycosylation, oxidation, adducts, terminal processing, or other detectable mass shifts.
Rapid Lot-to-Lot Comparison
Compare intact-mass profiles across production batches, candidates, stress conditions, or reference materials.
Reduced and Deglycosylated Analysis
Simplify complex spectra through optional reduction or enzymatic deglycosylation to evaluate chain masses and protein-backbone consistency.
ADC Drug-Load Profiling
Assess intact ADC mass, average drug-to-antibody ratio, and drug-load distribution using an approach selected for the conjugate chemistry.
Intact Mass Analysis Across Biologic Modalities
Sample preparation, chromatography, ionization conditions, and deconvolution parameters are tailored to the molecular architecture and expected heterogeneity of each modality.
- Monoclonal Antibodies and Fragments
Confirm intact mass, evaluate reduced heavy and light chains, and profile major glycoforms or terminal variants. - Antibody-Drug Conjugates
Characterize drug loading, average DAR, drug-load distribution, and major conjugate-related mass variants. - Recombinant Proteins and Enzymes
Verify expressed molecular mass and screen for truncation, processing variants, unexpected adducts, or major modifications. - Fusion Proteins and Multispecifics
Evaluate assembly, subunit composition, linker-related clipping, and complex proteoform patterns using tailored intact or subunit workflows. - Nanobodies and Protein Fragments
Obtain direct mass confirmation for smaller biologic formats with high resolving power and simplified spectral interpretation. - Biosimilars and Candidate Panels
Support rapid screening of mass consistency and major proteoform distributions across candidates, batches, or reference materials.
Analytical Approaches for Intact Mass Characterization
The appropriate strategy depends on the sample, expected heterogeneity, and the level of structural simplification required for confident interpretation.
| Approach | Best suited for | Typical information delivered |
| Denaturing intact LC-MS | Routine molecular-weight confirmation and rapid proteoform screening | Observed intact mass, theoretical-mass comparison, major mass shifts, and deconvoluted proteoform profiles. |
| Native mass spectrometry | Folded proteins, non-covalent assemblies, complex glycoforms, and suitable ADCs | Near-native intact masses, assembly state, major heterogeneity patterns, and project-dependent drug-load distributions. |
| Reduced or subunit analysis | Antibody chains, fragments, multispecifics, and complex intact spectra | Heavy- and light-chain masses or defined subunit masses that help resolve the source of whole-molecule heterogeneity. |
| Deglycosylated mass analysis | Glycoproteins whose glycan heterogeneity obscures the protein backbone | Simplified intact or subunit profiles supporting backbone-mass confirmation and differentiation of glycan-related from non-glycan mass shifts. |
Service Advantages
Our intact mass service combines rapid whole-molecule measurement with flexible sample preparation and expert interpretation.
- Complete Molecule View
Evaluate the undigested protein in a single measurement for direct top-level confirmation of molecular mass and major variants. - High Mass Accuracy
Use high-resolution accurate-mass instrumentation to distinguish meaningful mass differences in proteins and biotherapeutics. - Rapid Turnaround
Avoid routine enzymatic digestion and lengthy peptide-level processing when the project question can be answered at the intact level. - Fewer Preparation Artifacts
Reduce the risk of digestion-associated artifacts by analyzing the complete molecule with a streamlined preparation workflow. - Customizable Workflows
Add reduction, enzymatic deglycosylation, desalting, or modality-specific conditions according to the analytical objective. - Expert Data Interpretation
Receive scientist-reviewed raw and deconvoluted spectra, mass comparisons, and clear interpretation of major observed proteoforms.
Instrument Platform for Intact Mass Analysis
Creative Proteomics uses UHPLC coupled with high-resolution accurate-mass spectrometry, including Orbitrap- or Q-TOF-based platforms selected according to molecular size, heterogeneity, chromatographic behavior, and the required acquisition mode. Electrospray ionization generates a multi-charge envelope that is processed with mathematical deconvolution to determine the neutral molecular mass.
Platform configuration and data-processing parameters are optimized for the specific sample. Optional native, reduced, deglycosylated, or subunit workflows can be incorporated to simplify complex spectra and improve confidence in proteoform assignment.
Intact Mass Analysis Workflow
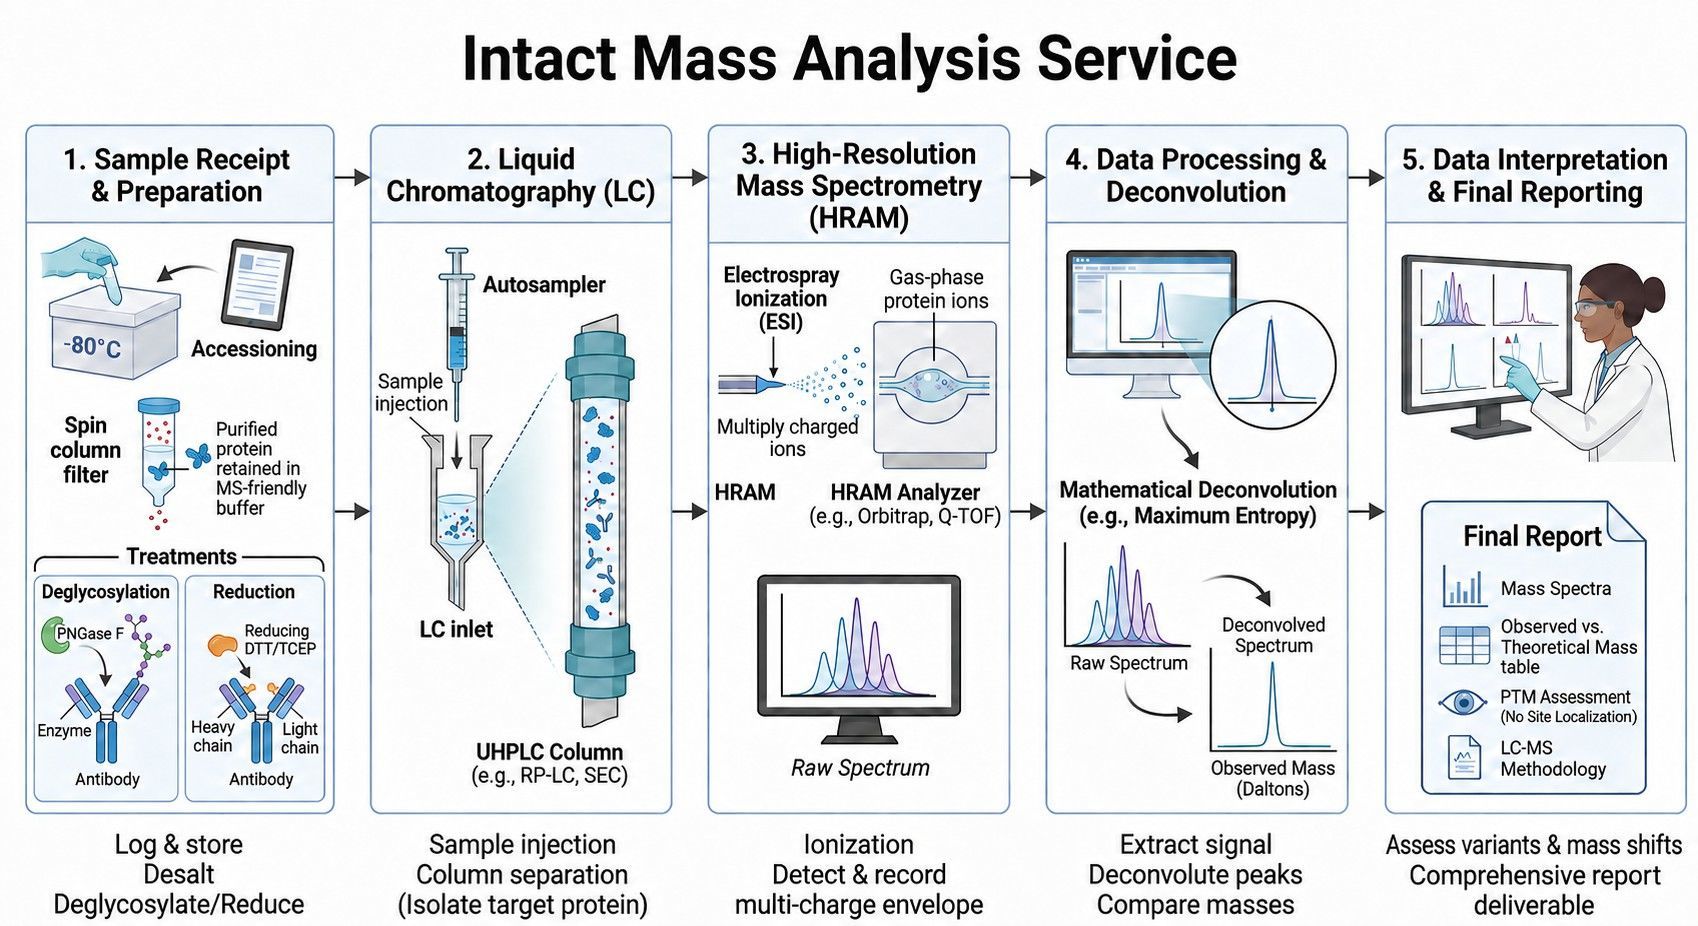
Our five-stage workflow covers sample receipt, LC-MS analysis, data processing, interpretation, and final reporting:
- 1
Sample Receipt and Preparation
Accession the sample, review concentration and buffer information, and perform fit-for-purpose desalting, buffer exchange, reduction, or deglycosylation as needed.
- 2
Liquid Chromatography
Introduce the prepared sample through an LC method selected to isolate the target protein and reduce matrix interference before mass detection.
- 3
High-Resolution Mass Spectrometry
Ionize the intact protein by electrospray and acquire its multi-charge envelope using high-resolution accurate-mass instrumentation.
- 4
Data Processing and Deconvolution
Convert the raw mass-to-charge spectrum into a neutral molecular-mass profile and extract observed peaks for comparison.
- 5
Data Interpretation and Final Reporting
Compare observed and theoretical masses, evaluate major variants and PTM-associated mass shifts, and prepare the final data package.
Report Deliverables
The standard intact mass data package includes:
- Raw Files: Secure delivery of native instrument data files for your internal archives.
- Executive Summary: Clear confirmation of whether the observed mass matches the theoretical mass.
- Methodology: Full documentation of LC-MS parameters and sample prep for reproducibility.
- Visuals: Publication-ready figures of both raw multi-charge and deconvolved mass spectrum.
Frequently Asked Questions About Intact Mass Analysis
What is the typical turnaround time?
Standard turnaround is 2-3 weeks from sample receipt. Expedited services are available upon request.
What types of samples can you analyze?
We routinely analyze a wide variety of proteins, including monoclonal antibodies (mAbs), antibody-drug conjugates (ADCs), recombinant proteins, nanobodies, and other biotherapeutics.
What buffer should my samples be submitted in?
Samples in PBS, Tris, HEPES, and other standard biological buffers may be accepted. Because nonvolatile salts are not directly MS-compatible, desalting or buffer exchange is performed before LC-MS analysis. Volatile, low-salt buffers are preferred when feasible.
What mass accuracy can I expect?
For many proteins and antibodies, high-resolution mass spectrometry can achieve mass accuracy within approximately 10-50 ppm. Actual performance depends on molecular size, heterogeneity, signal quality, calibration, and the selected analytical format.
How do you handle Post-Translational Modifications (PTMs)?
Intact mass analysis is effective for global modification profiling and can reveal major mass shifts associated with glycosylation, oxidation, terminal processing, or unexpected attachments. It generally does not localize a PTM within the sequence; peptide mapping or targeted PTM analysis is recommended when site-level localization is required.
Sample Requirements for Intact Mass Analysis
Please contact our technical team for tailored submission guidance. General planning guidelines are summarized below.
| Parameter | General guideline | Additional notes |
| Acceptable sample types | Monoclonal antibodies and fragments; ADCs; recombinant proteins; bispecific or multispecific antibodies; fusion proteins | Suitable for intact mass confirmation, drug-load profiling, and major structural-variant analysis. |
| Target concentration | Approximately 0.5-2.0 mg/mL | Final concentration requirements depend on molecular size, purity, and ionization response. |
| Minimum volume | Approximately 20 µL | Additional material is recommended when optional reduction, deglycosylation, or repeat analysis is requested. |
| Total protein amount | Approximately 1-10 µg analytical consumption | The submitted quantity may need to be higher; confirm the project-specific amount before shipment. |
| Recommended purity | Preferably >80% pure | Purity assessed by SDS-PAGE, SEC, or another suitable method supports signal-to-noise and reliable deconvolution. |
| Buffer composition | Volatile, MS-compatible buffers are preferred. | Standard biological buffers such as PBS, Tris, or HEPES may be accepted following project review and may require desalting or buffer exchange before analysis. |
Before shipping samples: Confirm sample identity, concentration, available volume, purity, buffer composition, storage conditions, expected molecular mass, and any requested reduction or deglycosylation treatment with our technical team.
References
- Wohlschlager T, Scheffler K, Forstenlehner IC, et al. Native mass spectrometry combined with enzymatic dissection unravels glycoform heterogeneity of biopharmaceuticals. Nature Communications. 2018;9:1713. doi:10.1038/s41467-018-04061-7.
- Debaene F, Boeuf A, Wagner-Rousset E, et al. Innovative native MS methodologies for antibody drug conjugate characterization: high resolution native MS and IM-MS for average DAR and DAR distribution assessment. Analytical Chemistry. 2014;86:10674-10683. doi:10.1021/ac502593n.
- Rosati S, Yang Y, Barendregt A, Heck AJR. Detailed mass analysis of structural heterogeneity in monoclonal antibodies using native mass spectrometry. Nature Protocols. 2014;9:967-976. doi:10.1038/nprot.2014.057.
- Yang Y, Liu F, Franc V, et al. Hybrid mass spectrometry approaches in glycoprotein analysis and their usage in scoring biosimilarity. Nature Communications. 2016;7:13397. doi:10.1038/ncomms13397.