Peptide Mapping Service
As biotherapeutics and novel proteins grow in complexity, thorough molecular characterization is critical during discovery and development. As a cornerstone of our Protein and Antibody Characterization portfolio, our high-resolution Peptide Mapping service provides detailed evidence supporting a biologic's identity, primary-sequence integrity, and selected product quality attributes.
Whether you are screening novel monoclonal antibodies in early discovery, performing clone selection, or optimizing upstream R&D processes, our team delivers molecular insights that help you advance your research with confidence.
- Confirm the primary amino-acid sequence with high-confidence LC-MS/MS evidence.
- Detect and localize sequence variants, clipping events, and selected PTMs.
- Compare candidates, clones, batches, formulations, and stress conditions.
In line with ICH Q6B Guidance for biologic characterization
Jump to Section
- Why Peptide Mapping Matters
- Service Capabilities
- Biologic Modalities
- Analytical Approaches
- Service Advantages
- Instrument Platform
- Workflow
- Report Deliverables
- FAQs
- Sample Requirements
Why Peptide Mapping Matters in Biologic Development
Peptide mapping is the fingerprinting of a protein at the peptide level. Because intact proteins are often too large and heterogeneous for detailed amino-acid-level analysis, the target is digested into smaller peptides using one or more sequence-specific proteases.
The resulting peptides are separated and analyzed by high-resolution liquid chromatography-tandem mass spectrometry. Their accurate masses, retention behavior, and fragment-ion spectra create a characteristic map that can be compared with a theoretical sequence, candidate molecule, production clone, or reference sample.
This bottom-up view supports primary-sequence confirmation and can reveal unintended substitutions, truncation or clipping, oxidation, deamidation, glycosylation, and other project-relevant modifications. When suitable methods and controls are used, selected modifications can also be localized and compared across samples.
Peptide mapping is therefore valuable for candidate screening, clone selection, process research, comparability, formulation studies, and forced-degradation investigations. It also complements intact mass analysis, which provides the whole-molecule view but generally cannot localize a mass difference within the protein sequence.
Peptide Mapping Service Capabilities
Our LC-MS/MS peptide mapping platform supports the following core characterization needs:
Primary Sequence Confirmation
Match identified peptides and fragment-ion evidence against the expected protein sequence to establish a high-confidence peptide map.
Sequence Coverage Assessment
Calculate overall and region-specific coverage and identify technically challenging or unobserved portions of the sequence.
PTM Identification and Localization
Detect and localize selected modifications such as oxidation, deamidation, glycosylation, phosphorylation, or terminal processing when supported by the data.
Mutation and Variant Analysis
Investigate unintended amino-acid substitutions, truncations, clipping events, or other sequence differences in candidate molecules.
Relative Modification Comparison
Compare the relative abundance of selected modified and unmodified peptides across samples, treatments, batches, or stress conditions.
Specialized Structural Mapping
Add non-reduced digestion, deglycosylation, or tailored fragmentation workflows for disulfide-linked peptides and other project-specific targets.
Peptide Mapping Across Biologic Modalities
Digestion strategy, LC separation, fragmentation, and data analysis are tailored to the molecular architecture and expected modifications of each biologic format.
- Monoclonal Antibodies
Confirm heavy- and light-chain sequence coverage and characterize common antibody modifications, terminal variants, glycopeptides, or degradation hotspots. - Bispecific and Multispecific Antibodies
Use chain-aware sequence databases and complementary digestion strategies to distinguish related domains, chains, and engineered regions. - Antibody-Drug Conjugates
Map the antibody sequence and, when analytically feasible, investigate conjugation-related peptides or payload-associated changes. - Fusion and Glycosylated Proteins
Evaluate fusion junctions, linkers, processing sites, and glycopeptides using optional deglycosylation or complementary proteases. - Recombinant Proteins and Enzymes
Verify the expressed sequence and assess mutations, clipping, oxidation, deamidation, or other project-relevant modifications. - Candidate and Clone Panels
Compare structural fidelity across expression clones, culture conditions, process variables, or early-stage candidate molecules.
Analytical Approaches for Peptide Mapping
The final workflow is selected according to the desired sequence coverage, modification targets, sample complexity, and comparison objective.
| Approach | Best suited for | Typical information delivered |
| Standard tryptic mapping | Routine sequence confirmation and broad PTM screening | Tryptic peptide identifications, sequence coverage, selected PTMs, annotated MS/MS spectra, and peptide-level comparison. |
| Multi-enzyme mapping | Coverage gaps, hydrophobic regions, long or short tryptic peptides, and complex constructs | Complementary peptide sets generated with enzymes such as chymotrypsin, Glu-C, or Asp-N to improve coverage and assignment confidence. |
| PTM-focused mapping | Known modification hotspots, forced-degradation studies, and formulation comparisons | Targeted or expanded searches, site localization, and project-dependent relative comparison of modified and unmodified peptides. |
| Non-reduced or specialized mapping | Disulfide connectivity, scrambled disulfides, glycopeptides, or difficult sequence variants | Project-specific digestion and fragmentation designed to preserve linked peptides or obtain diagnostic fragment ions. |
Service Advantages
Our service combines advanced LC-MS/MS technology, flexible digestion strategies, and scientist-led interpretation.
- High Sequence Coverage
Apply optimized preparation and complementary proteases to maximize sequence coverage, including technically challenging regions. - Evidence-Based PTM Localization
Use high-resolution MS/MS evidence to localize selected modifications to specific peptides and, when fragment-ion coverage is sufficient, individual residues. - Customized Workflows
Tailor reduction, alkylation, digestion, deglycosylation, chromatography, and fragmentation to the molecule and research question. - Integrated Characterization
Combine peptide mapping with intact mass, glycan profiling, free-thiol, disulfide, or aggregation analysis for a broader molecular picture. - Advanced Instrumentation
Use high-resolution Orbitrap- or timsTOF-based LC-MS/MS platforms selected for coverage, sensitivity, and fragmentation needs. - Expert Scientific Support
Receive project consultation, method selection, data review, interpretation, and post-report technical support from experienced scientists.
Instrument Platform for Peptide Mapping
Creative Proteomics uses high-resolution LC-MS/MS platforms, including Thermo Orbitrap and Bruker timsTOF systems, coupled with reversed-phase UHPLC separation and experiment-specific fragmentation methods. Platform selection depends on peptide complexity, modification targets, coverage requirements, and whether specialized fragmentation is needed.
Data processing combines accurate-mass precursor information, MS/MS fragment-ion matching, sequence-database searching, and manual review of project-critical peptides. Search parameters and modification lists are defined according to the study objective to balance discovery depth with assignment confidence.
Peptide Mapping Workflow
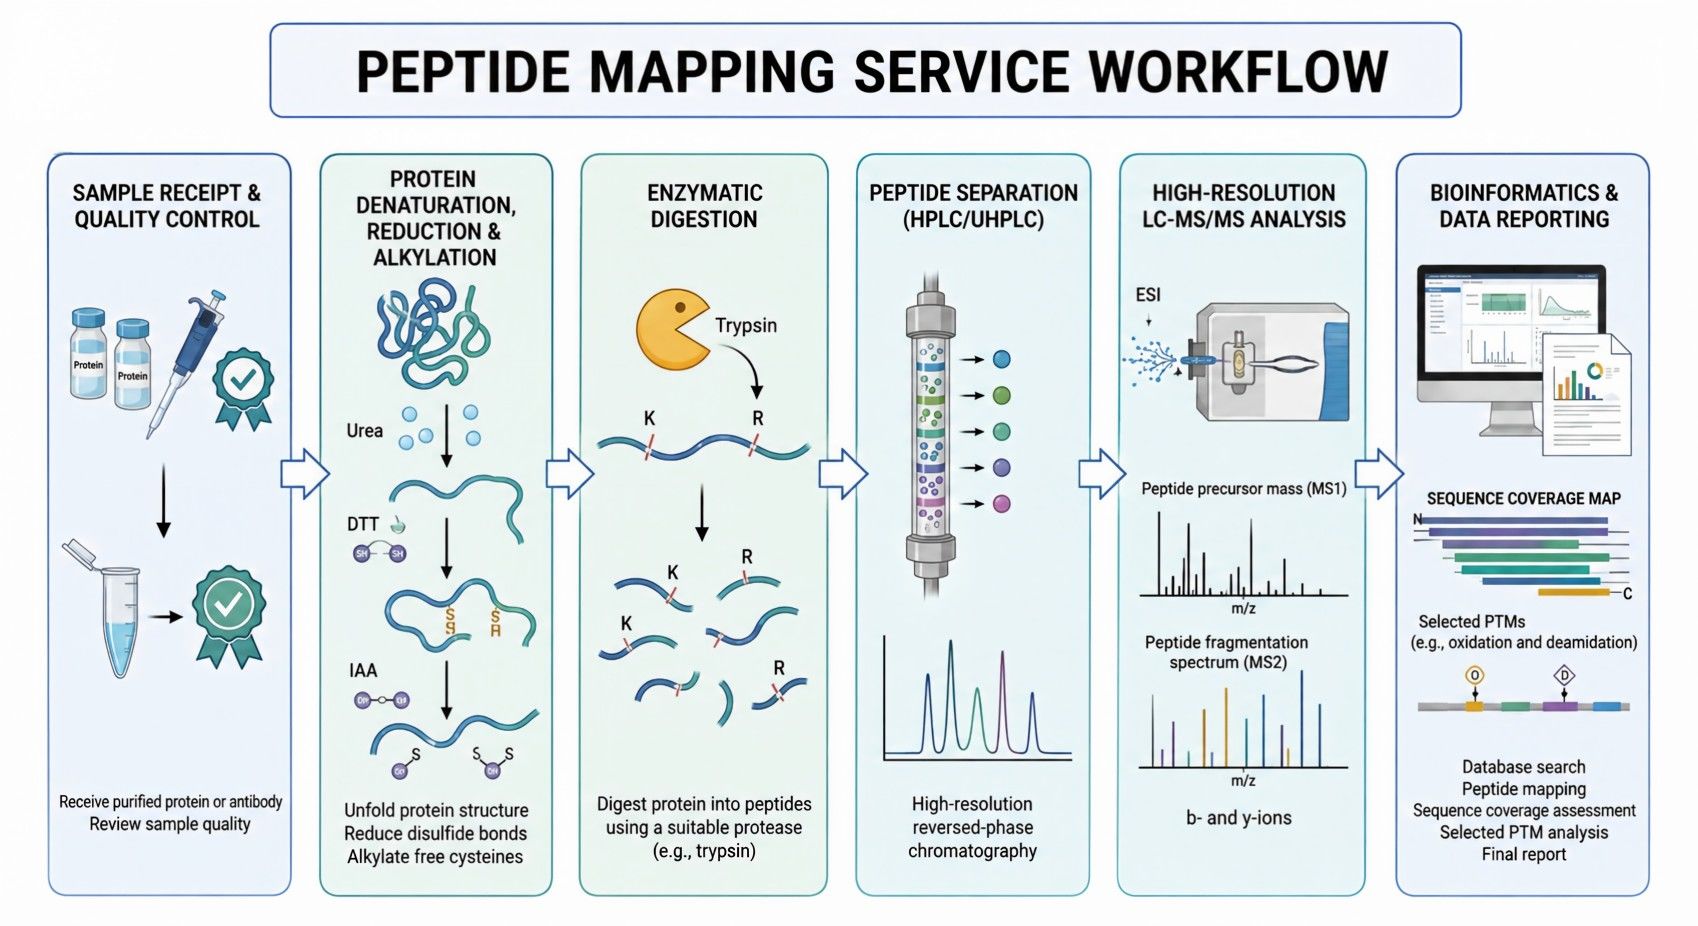
Our peptide mapping workflow consists of six sequential stages:
- 1
Sample Receipt and Quality Control
We accession the sample and review its identity, concentration, purity, buffer composition, storage conditions, and project objectives to confirm suitability for analysis.
- 2
Protein Denaturation, Reduction, and Alkylation
The protein is denatured to expose digestion sites. For a standard reduced workflow, disulfide bonds are reduced and free cysteines are alkylated under controlled conditions.
- 3
Enzymatic Digestion
The protein is digested with trypsin or another suitable protease selected according to the sequence, coverage requirements, and analytical objectives.
- 4
Peptide Separation
The resulting peptides are separated by high-resolution reversed-phase HPLC or UHPLC to reduce spectral complexity and improve peptide detection.
- 5
High-Resolution LC-MS/MS Analysis
Accurate peptide precursor masses and MS/MS fragment-ion spectra are acquired using project-appropriate ionization and fragmentation conditions.
- 6
Bioinformatics and Data Reporting
The spectra are searched against the expected protein sequence to generate peptide identifications and sequence coverage maps. Selected PTMs and sequence variants are evaluated when supported by the data, and relative comparisons are performed where appropriate. The results are reviewed and compiled into the final report.
Complementary digestion. For complex proteins such as monoclonal antibodies, auxiliary digestion with chymotrypsin, Glu-C, Asp-N, or another suitable protease may improve coverage of regions that are not well represented by trypsin alone.
Report Deliverables
Upon completion, you will receive a research report tailored to your project goals, including:
- Comprehensive summary: an overview of the methodology, analytical approach, and key findings.
- Sequence coverage maps: visual maps showing total and region-specific amino-acid coverage.
- Selected PTM analysis: Identification and evidence-based localization of selected modifications, with project-dependent relative comparison across samples where appropriate.
- Annotated spectra: MS/MS evidence for key peptides, sequence variants, or modifications.
- Raw data: secure transfer of raw mass-spectrometry files for internal archives or bioinformatics workflows.
Frequently Asked Questions About Peptide Mapping
Can you map complex, heavily glycosylated proteins or fusion proteins?
For heavily glycosylated proteins, deglycosylation may be used to improve protein-backbone sequence coverage, while glycopeptide-focused workflows are required when site-specific glycosylation information is needed.
Can you identify, localize, and compare post-translational modifications (PTMs)?
Yes. High-resolution peptide mapping can identify and localize selected PTMs when supported by sufficient MS/MS evidence. For suitable modified and unmodified peptides and an appropriate study design, we can also compare their relative abundance across samples. Feasibility depends on the modification type, peptide properties, sample quality, and analytical method.
Can your analysis differentiate between Leucine (Leu) and Isoleucine (Ile)?
Leucine and isoleucine are isobaric and generally cannot be distinguished by MS1 or routine MS/MS peptide mapping. In selected cases, differentiation may be investigated using specialized fragmentation, multistage MS, differential enzymatic digestion, or other orthogonal evidence. Feasibility should be evaluated for the specific target peptide.
What is the difference between peptide mapping and intact mass analysis?
Intact mass analysis measures the mass of the whole protein and provides an overall view of molecular mass and heterogeneity, but it generally cannot determine where a mass difference occurs. Peptide mapping digests the protein into smaller peptides and provides peptide-level information for sequence confirmation, selected PTMs, and sequence variants. Residue-level localization may be achieved when supported by sufficient diagnostic fragment-ion evidence.
Sample Requirements for Peptide Mapping
Please contact our technical team for tailored submission guidance. General sample requirements are summarized below.
| Parameter | General guideline | Additional notes |
| Protein amount | Preferably 50-100 µg | Lower-input feasibility may be assessed depending on molecular size, purity, complexity, and project scope. |
| Concentration | Preferably at least 1.0 mg/mL | Higher concentration may support recovery during desalting and multiple digestion or repeat-analysis conditions. |
| Buffer | Volatile or MS-compatible buffers are preferred; PBS may be accepted | Provide the complete formulation. Desalting or buffer exchange can be included when needed. |
| Components to avoid | High Tris, glycerol, PEG, SDS, Triton X-100, or other strong detergents | These components may interfere with digestion, chromatography, ionization, or data interpretation. |
| Shipping | Ship frozen on dry ice in tightly sealed, clearly labeled tubes | Use low-binding containers where practical and include the completed Sample Submission Form. |
Before shipping samples, please confirm the sample identity, expected sequence, concentration, amount, purity, buffer composition, storage conditions, comparison groups, and any priority PTMs or sequence regions with our technical team.
References
- Mouchahoir T, Schiel JE. Development of an LC-MS/MS peptide mapping protocol for the NISTmAb. Analytical and Bioanalytical Chemistry. 2018;410:2111-2126. doi:10.1007/s00216-018-0848-6.
- Rogers RS, Nightlinger NS, Livingston B, et al. Development of a quantitative mass spectrometry multi-attribute method for characterization, quality control testing and disposition of biologics. mAbs. 2015;7:881-890. doi:10.1080/19420862.2015.1069454.
- Jiang P, Li F, Ding J. Development of an efficient LC-MS peptide mapping method using accelerated sample preparation for monoclonal antibodies. Journal of Chromatography B. 2020;1137:121895. doi:10.1016/j.jchromb.2019.121895.