Mass Spectrometry-Based Drug Target Identification & Deconvolution
Target deconvolution is the systematic identification of cellular protein targets through which a bioactive small molecule, natural product, or therapeutic lead exerts its phenotypic effect. Chemical proteomics couples biophysical or chemical target capture with high-resolution quantitative mass spectrometry (LC-MS/MS) to measure drug-protein engagement across the entire native proteome.
Unlike classical affinity approaches that require a derivatized ligand, modern chemoproteomics offers both label-free and probe-based modalities. Depending on the method, the resulting evidence may include thermal stability shifts, dose-dependent protease protection or competition trends, probe-enrichment changes, and peptide-level structural responses. These readouts support target prioritization, lead optimization, and research-stage safety investigation; dedicated biophysical assays are used when equilibrium binding constants are required.
Content Guide
- When to Apply Chemoproteomics
- Method Selection Matrix
- What Problems We Solve
- Service Advantages
- Core Platforms & Technologies
- Step-by-Step Workflow
- Sample Requirements & Prep
- Deliverables & Data Packages
When to Apply Chemoproteomics Target Deconvolution
- Phenotypic Hit Deconvolution: Identify direct protein targets of bioactive compounds identified from cell-based phenotypic screens.
- Natural Product & Metabolite Target Discovery: Unravel direct cellular receptors of unmodified botanical extracts, secondary metabolites, and endogenous ligands.
- Off-Target & Safety Liability Profiling: Screen drug candidates for secondary target-engagement signals that may help explain polypharmacology or potential safety liabilities.
- Mechanism of Action (MoA) Elucidation: Differentiate physical target engagement from downstream signaling network rewiring and secondary effector cascades.
- PROTAC & Molecular Glue Target Mapping: Characterize ternary complex formation between E3 ligases, small-molecule degraders, and neo-substrates.
- Drug Repurposing Research: Generate new target hypotheses and mechanism evidence for established compounds in research models.
Chemoproteomics Method Selection & Comparison Matrix
Match your chemical entity and study design with the optimal target deconvolution strategy.
| Method | Chemical Modification | Matrix Compatibility | Binding Mechanism | Structural Resolution | Optimal Compound Class |
|---|---|---|---|---|---|
| Thermal Proteome Profiling (TPP / CETSA) | None (Label-Free) | Living cells, lysates, tissues | Non-covalent & covalent | Whole protein | Synthetic drugs, natural products, clinical leads |
| Drug Affinity Responsive Target Stability (DARTS) | None (Label-Free) | Native cell lysates, tissue extracts | Non-covalent & covalent | Whole protein / domain | Unmodified small molecules, fragile compounds |
| Limited Proteolysis (LiP-MS) | None (Label-Free) | Complex cell lysates, organelles | Non-covalent & covalent | Peptide-level structural region | All small molecules, metabolites, allosteric leads |
| Activity-Based Protein Profiling (ABPP) | Probe required (Electrophile + Tag) | Living cells, lysates, membranes | Covalent / Mechanism-based | Protein / probe-labeled site | Covalent inhibitors, kinase/protease inhibitors |
| Photoaffinity Labeling (PAL) & Capture | Probe required (Photophore + Bioorthogonal Tag) | Living cells, lysates | Non-covalent → Covalent crosslink | Protein & crosslinked peptide | Low-affinity binders, reversible ligands, lipid probes |
What Problems Does Chemical Proteomics Solve?
Overcoming the critical blind spots of recombinant assays and transcriptomic profiling.
Unmodified Compound Deconvolution
Deconvolute fragile natural products and unmodified lead molecules via TPP, DARTS, and LiP-MS without chemical tag synthesis.
Native Complex & Membrane Targets
Capture drug-target engagement in intact living cells, preserving multi-protein complexes, endogenous cofactors, and GPCR conformations.
Physical Binding vs. Downstream Effects
Prioritize target-engagement evidence from TPP, LiP-MS, ABPP, or PAL separately from downstream abundance and pathway responses.
Whole-Proteome Off-Target Profiling
Profile proteome-scale target-engagement patterns to identify candidate secondary targets and potential polypharmacology for follow-up.
Binding Pocket & Allosteric Localization
Use LiP-MS to localize ligand-responsive peptide regions and structural changes that can guide binding-site hypotheses and orthogonal follow-up.
Complex and Low-Abundance Target Contexts
Method selection can combine enrichment, fractionation, DIA, or ion-mobility-enabled acquisition when matrix complexity or low target abundance makes confident quantification more difficult.
Advantages of Our Drug Target Identification Services
Multi-Platform Flexibility
Label-Free & Probe-Based
Comprehensive portfolio spanning TPP, DARTS, LiP-MS, ABPP, and photoaffinity capture under one roof.
Deep Proteome Coverage
Proteome-Scale Readout
High-resolution quantitative mass spectrometry supports broad target-engagement profiling, with acquisition depth adapted to matrix, sample amount, and study design.
Physiological Relevance
Living Cells & Tissues
Target-deconvolution strategies can be adapted to intact cells, primary cells, tissue specimens, lysates, and selected subcellular contexts according to method compatibility.
Binding Site Resolution
Peptide-Level Mapping
Structural footprinting via LiP-MS identifies ligand-responsive peptide regions and conformational changes that can refine binding-site hypotheses.
Multi-Tiered QC Strategy
Specificity-Focused Controls
Matched controls, concentration-response patterns, inactive analogs when available, and competition experiments help reduce non-specific target candidates.
Validation Continuity
PRM & CETSA Follow-Up
Prioritized candidates can move into targeted MS, cellular CETSA, biophysical binding assays, or functional experiments according to the validation question.
Core Chemoproteomics Platforms & Technologies
Integrated biophysical and chemical biology platforms for comprehensive target deconvolution
Thermal Proteome Profiling (TPP / CETSA-MS)
- Measures compound-associated changes in protein thermal stability across a project-specific temperature design.
- Temperature-range TPP and concentration-dependent thermal profiling can be selected according to the target-deconvolution question.
- Quantitative MS strategy: TMT-based multiplexing remains established for TPP, while label-free DIA can provide an alternative route for thermal-shift quantification.
Drug Affinity Responsive Target Stability (DARTS)
- Tests whether ligand exposure changes susceptibility of proteins to controlled limited proteolysis in native lysates.
- Useful for unmodified compounds when chemical tagging is undesirable, with candidate confidence strengthened by concentration-response and orthogonal evidence.
- Quantitative MS strategy: Broad LC-MS/MS readout is selected to balance proteome coverage, reproducibility, and sample throughput.
Limited Proteolysis-Coupled MS (LiP-MS)
- Generates structurally informative cleavage patterns in complex native lysates and compares ligand-treated versus control conditions.
- Identifies ligand-responsive peptide regions and conformational changes, which can approximate binding regions without assuming single-residue resolution.
- Quantitative MS strategy: DIA-based LiP-MS can improve peptide-level reproducibility and data continuity in complex semitryptic/tryptic datasets.
Activity-Based Protein Profiling (ABPP) & PAL
- Uses reactive or photoaffinity probes to enrich proteins or sites that can be interrogated competitively with an unlabeled compound.
- Concentration-series designs can generate assay-dependent competition curves and site/protein engagement metrics rather than a universal equilibrium binding constant.
- Quantitative MS strategy: DIA can support reproducible site-centric or enriched chemoproteomic profiling across larger condition sets.
How DIA and 4D-DIA Support Drug Target Deconvolution
DIA and 4D-DIA are quantitative mass-spectrometry acquisition strategies, not standalone target-engagement assays. Their role is to improve the consistency and depth of the proteomic readout generated by TPP, LiP-MS, ABPP/PAL, or global mechanism-of-action studies. The target-identification logic still comes from the experimental perturbation and controls.
| Target-Deconvolution Context | Role of DIA / 4D-DIA | Interpretation Boundary |
|---|---|---|
| TPP / CETSA-MS | DIA quantitative proteomics can provide label-free thermal-profile quantification when the study benefits from a non-isobaric acquisition strategy. | Thermal stability changes support target-engagement hypotheses but may also reflect indirect protein-state changes; orthogonal confirmation can be important. |
| LiP-MS | DIA supports reproducible quantification of structurally informative peptide features. When additional gas-phase separation is useful, 4D-DIA quantitative proteomics can be considered for complex peptide mixtures. | LiP-MS identifies ligand-responsive peptide regions and conformational changes; it does not universally assign a single binding residue. |
| ABPP / PAL | DIA can scale site-centric or enrichment-based competition profiling across larger compound, concentration, or time-course designs. | Competition curves are assay-dependent engagement readouts and should not be presented automatically as equilibrium affinity constants. |
| Global MoA Proteomics | Discovery proteomics using DIA or 4D-DIA can map treatment-associated abundance changes and pathway responses after compound exposure. | Global abundance changes describe downstream response and do not by themselves prove direct physical binding. |
After candidate targets are prioritized, targeted proteomics can support focused PRM/MRM follow-up in additional research samples, while CETSA, SPR, ITC, MST, or functional perturbation can answer complementary validation questions.
Step-by-Step Target Deconvolution Workflow
Our 5-step workflow connects study scoping, target-engagement experiments, quantitative MS, evidence-based prioritization, and an appropriate validation roadmap.
Evaluate compound structure, modifiability, matrix type, and mechanism to select the optimal label-free (TPP, DARTS, LiP-MS) or probe-based approach.
Incubate living cells or native lysates with compound concentration series alongside vehicle and competition controls. Apply thermal challenge, limited proteolysis, or UV crosslinking.
Select TMT-based multiplexing, DIA, or ion-mobility-enabled DIA/4D-DIA according to the assay format, matrix complexity, number of conditions, and quantitative objective.
Apply project-appropriate identification and statistical QC, thermal-curve fitting, concentration-response modeling, probe-enrichment analysis, or LiP-MS peptide-level structural analysis as required by the selected method.
Deliver ranked target candidates with method-specific engagement metrics, biological context, evidence limitations, and recommended targeted or orthogonal follow-up.
- Label-free and probe-based target deconvolution pipelines tailored to compound chemistry
- Target-engagement profiling in intact cells, primary cells, tissue specimens, and native lysates when method-compatible
- Multi-layered false-positive exclusion using vehicle subtraction and competition assays
- Research support from initial phenotypic hit through candidate prioritization and orthogonal target validation
Sample Requirements & Preparation Guidelines
Preserve Native Protein State: TPP, DARTS, and LiP-MS depend on controlled protein stability or conformation. Lysis and handling conditions should therefore be selected to preserve the native state required by the chosen assay.
Match the Vehicle: Provide the solvent or vehicle used for compound preparation, together with available solubility and stability information.
| Material | Planning Information | Main Considerations |
|---|---|---|
| Intact Cells | Cell type, treatment design, expected viability, number of conditions, and whether live-cell target engagement is essential | Cell permeability, treatment duration, thermal or probe compatibility, and biological replication |
| Cell or Tissue Lysates | Matrix, available total protein, lysis history, buffer composition, and storage conditions | Preservation of native structure, detergent compatibility, protease activity, and freeze-thaw history |
| Research Tissue Specimens | Tissue type, preservation method, available material, heterogeneity, and whether intact tissue or lysate analysis is intended | Sampling consistency, endogenous proteolysis, storage history, and extraction feasibility |
| Test Compound / Probe | Identity, available amount, purity information, stock concentration, vehicle, solubility, and active/inactive analogs when available | Number of concentrations, replicate structure, probe derivatization feasibility, and competition-control design |
*Final input requirements are method- and matrix-dependent. Share your available material and experimental design so the target-deconvolution workflow can be scoped before sample submission.
Data Analysis & Target Deconvolution Deliverables
Clear, auditable, and publication-ready target discovery reports
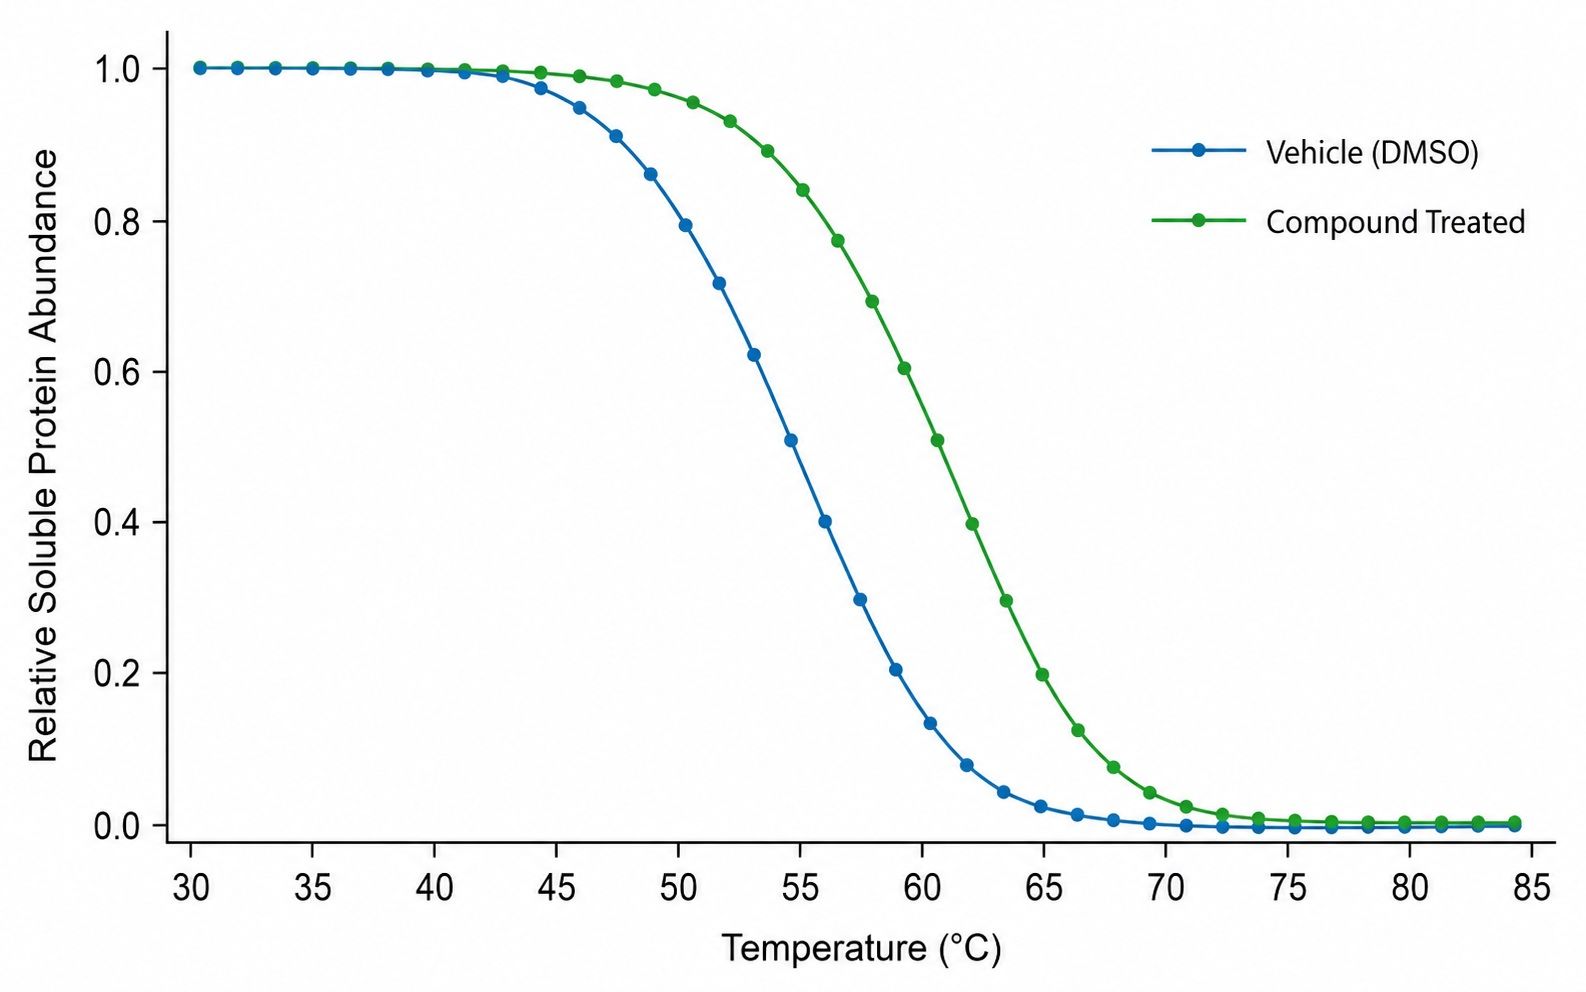
Thermal Melting Shift Curves: Representative vehicle-versus-compound thermal profiles showing calculated melting points (Tm) and compound-associated thermal shifts (ΔTm), including stabilization or destabilization where observed.
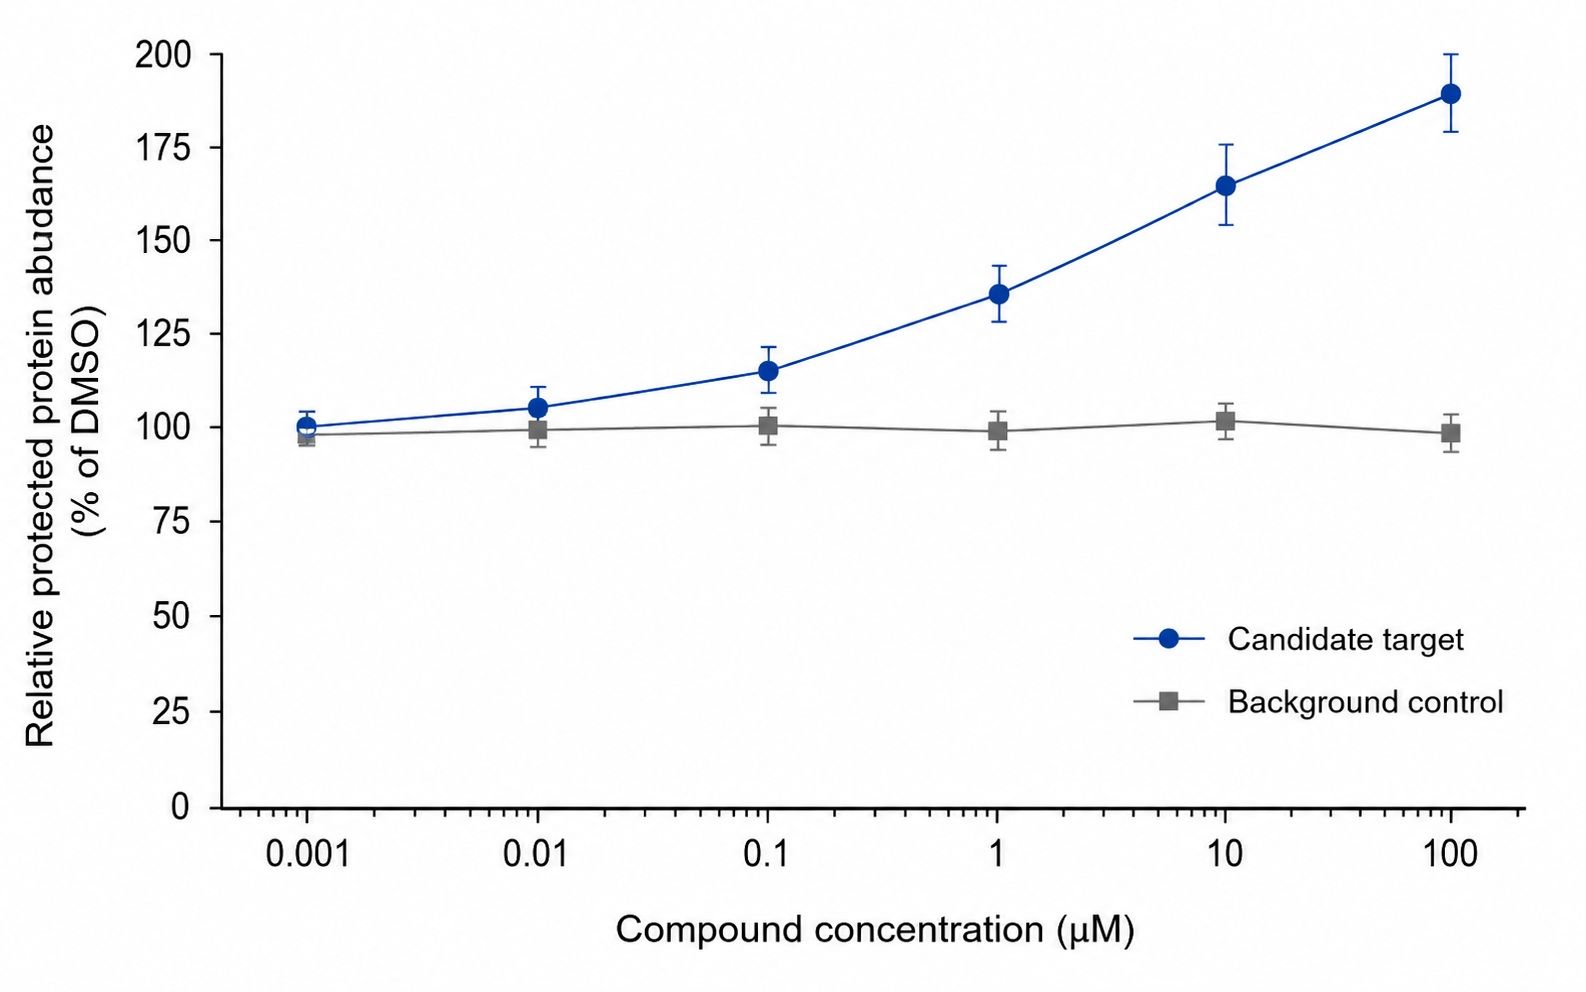
Dose-Dependent Protease Protection: Quantitative titration curves demonstrating ligand-induced proteolytic stabilization across concentration series in DARTS.
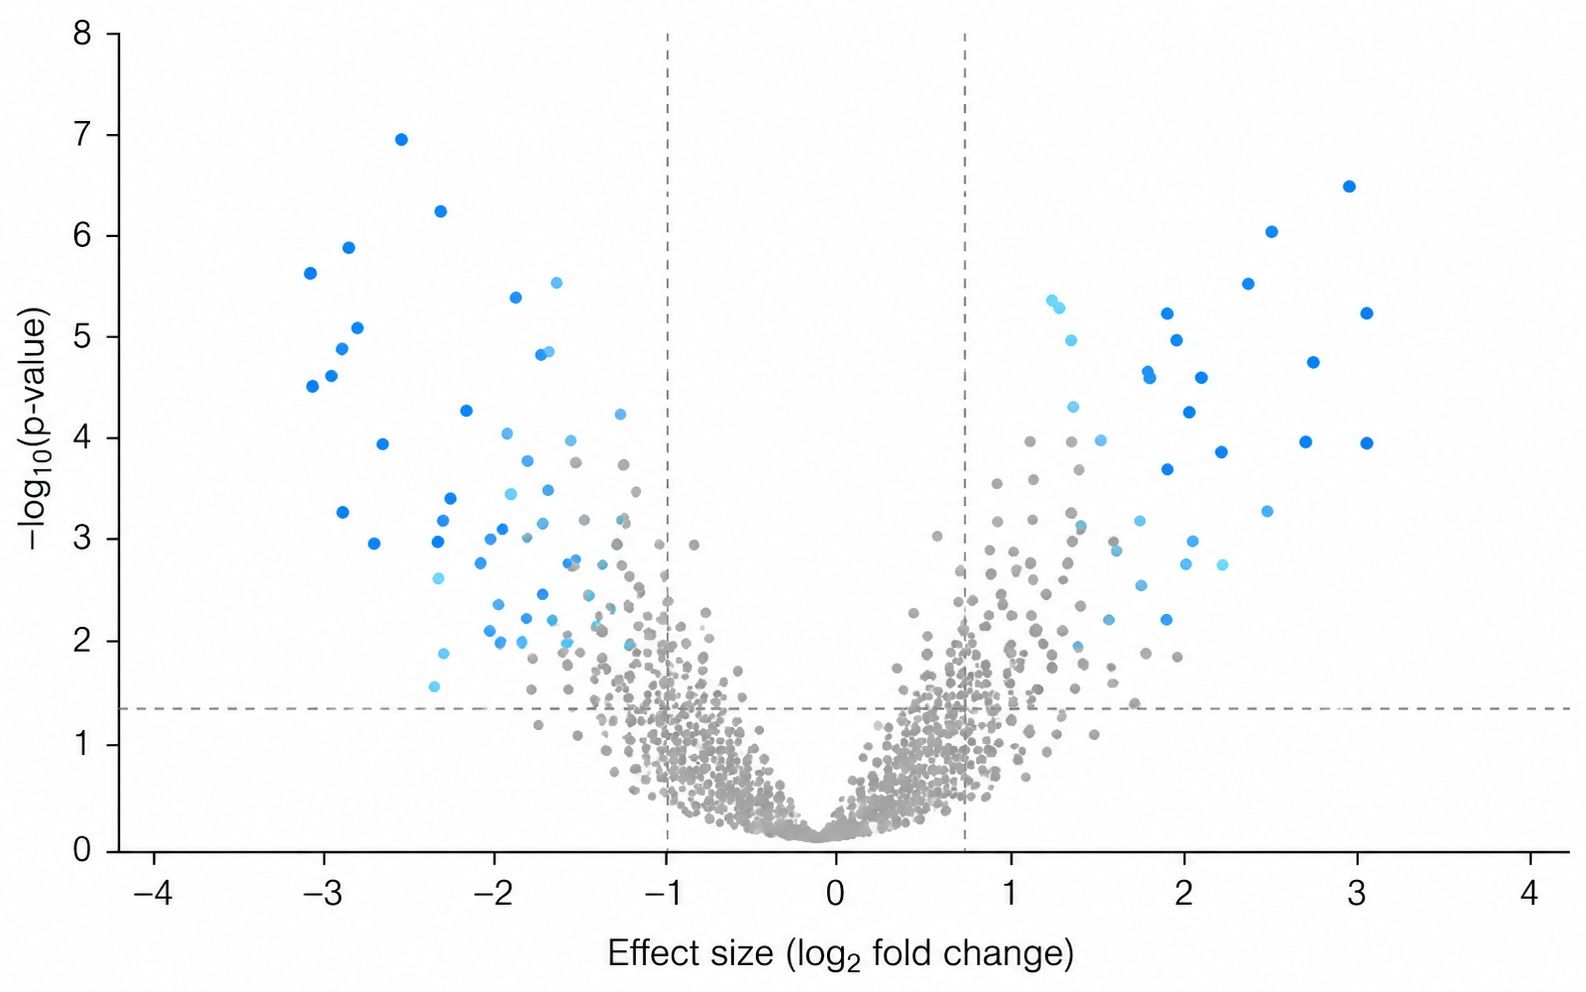
Target Candidate Volcano Plot: Statistical visualization of effect size and significance, highlighting candidate target-engagement signals for prioritization rather than treating every significant protein as a confirmed direct target.
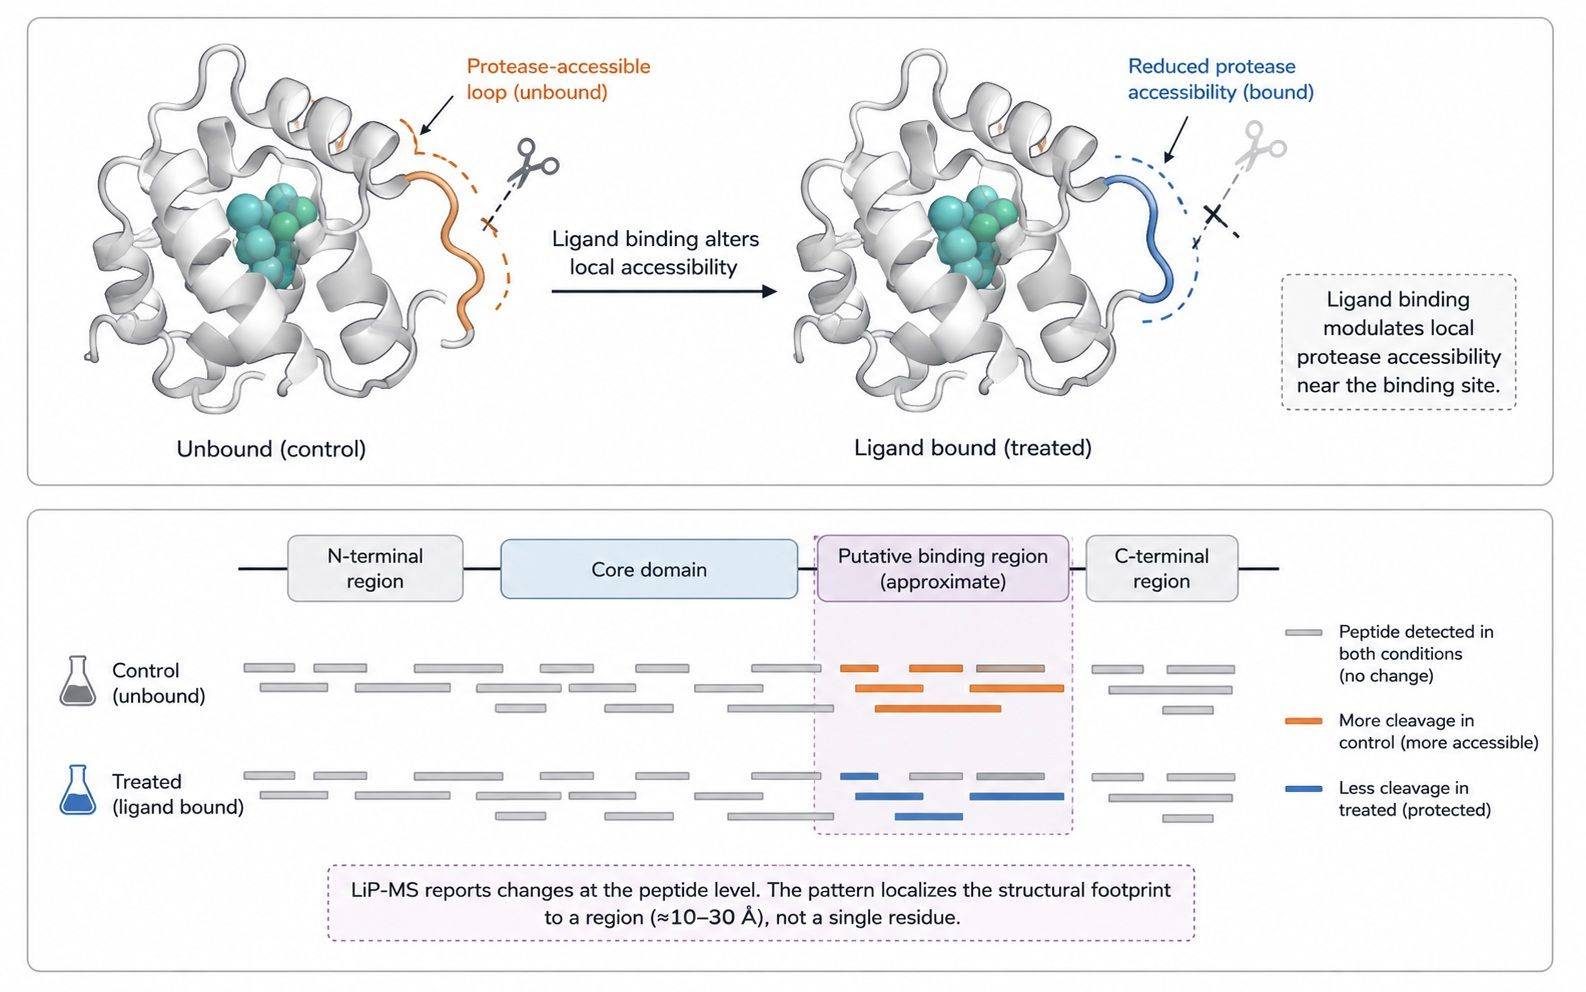
LiP-MS Structural Footprinting Map: Peptide-level coverage maps showing ligand-responsive structural regions that can guide binding-site hypotheses.
Standard Deliverables Checklist
- Raw mass spectrometry instrument files (.raw / .d) and search parameter logs
- Comprehensive target identification matrix with UniProt accessions and gene symbols
- Statistical ranking tables with method-appropriate fold changes, adjusted significance metrics, thermal shifts, protection scores, competition responses, or peptide-level structural evidence
- Method-specific concentration-response curves and thermal denaturation profiles when applicable
- Peptide-level structural footprinting and binding pocket maps (for LiP-MS)
- Biological pathway enrichment and Gene Ontology (GO) annotation
- Identification, quantification, replicate, and statistical quality-control summaries appropriate to the selected workflow
- Target validation roadmap with recommended targeted MS, CETSA, biophysical, or functional follow-up
Drug Target Identification Frequently Asked Questions
Representative Research Study: Thermal Proteome Profiling Maps Drug Targets in Living Cells
Tracking Cancer Drugs in Living Cells by Thermal Profiling of the Proteome
Journal: Science · Published: 2014 · DOI: 10.1126/science.1255784
7,000+
Proteins profiled in parallel
50+ Targets
Targets delineated for staurosporine
Intact Cells
Native physiological context
Unmodified Compound
No affinity-tag derivatization
Study Overview & Objectives
Researchers used Thermal Proteome Profiling (TPP) to determine thermal profiles for more than 7,000 proteins in human cells and to examine how small-molecule treatment altered protein thermal stability. The study included kinase inhibitors such as staurosporine and dasatinib and demonstrated that proteome-wide thermal shifts can reveal expected targets, additional targets, and downstream protein-state changes in intact cells.
Technical Strategy & Results
| Research Dimension | Experimental Approach | Critical Outcome |
|---|---|---|
| Intact Cellular Profiling | Living K562 cells incubated with kinase inhibitors across a temperature gradient (37°C–67°C) | Detected target-engagement and drug-responsive thermal shifts in an intact-cell context with endogenous cofactors and protein complexes |
| Quantitative Mass Spectrometry | Multiplexed TMT 10-plex isobaric labeling on high-resolution Orbitrap systems | Generated proteome-wide thermal profiles for more than 7,000 proteins across the study |
| Off-Target & Polypharmacology Mapping | Compared thermal shifts (ΔTm) between broad-spectrum and selective inhibitors | Revealed additional drug-responsive targets and thermal shifts in downstream pathway proteins; downstream shifts were not interpreted automatically as direct binding |
Key Findings & Process Implications
- Proteome-Wide Target Engagement: TPP detected known drug targets and additional thermal-response proteins without requiring an affinity-tagged compound.
- Direct and Indirect Thermal Responses: The study showed that thermal shifts can reflect direct ligand binding as well as downstream protein-state changes, reinforcing the need to interpret candidates in biological context.
- Target-Deconvolution Framework: The work established thermal profiling as a broadly useful route for unbiased target-engagement studies in living cells and helped define the modern TPP workflow.
Selected Scientific References:
- Savitski MM, Reinhard FBM, Franken H, et al. Tracking cancer drugs in living cells by thermal profiling of the proteome. Science. 2014;346(6205):1255784. doi:10.1126/science.1255784.
- Lomenick B, Hao R, Jonai N, et al. Target identification using drug affinity responsive target stability (DARTS). Proc Natl Acad Sci USA. 2009;106(51):21984-21989. doi:10.1073/pnas.0910040106.
- Feng Y, De Franceschi G, Kahraman A, et al. Global analysis of protein structural changes in complex systems. Nat Biotechnol. 2014;32(10):1036-1044. doi:10.1038/nbt.2999.
- Cravatt BF, Wright AT, Kozarich JW. Activity-based protein profiling: chemical approaches to explore genome biology. Annu Rev Biochem. 2008;77:383-414. doi:10.1146/annurev.biochem.75.101304.124125.
- George AL, Sidgwick FR, Watt JE, et al. Comparison of Quantitative Mass Spectrometric Methods for Drug Target Identification by Thermal Proteome Profiling. J Proteome Res. 2023;22(8):2629-2640. doi:10.1021/acs.jproteome.3c00111.
- Malinovska L, Cappelletti V, Kohler D, et al. Proteome-wide structural changes measured with limited proteolysis-mass spectrometry: an advanced protocol for high-throughput applications. Nat Protoc. 2023;18(3):659-682. doi:10.1038/s41596-022-00771-x.
- Yang F, Jia G, Guo J, Liu Y, Wang C. Quantitative Chemoproteomic Profiling with Data-Independent Acquisition-Based Mass Spectrometry. J Am Chem Soc. 2022;144(2):901-911. doi:10.1021/jacs.1c11053.